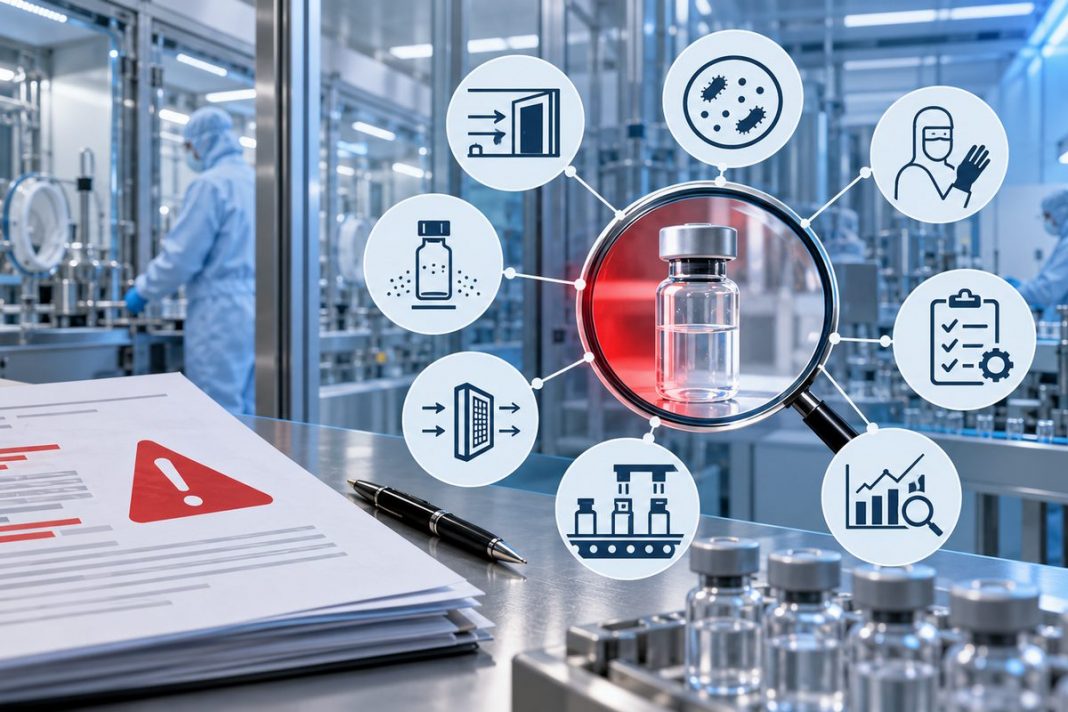
A September 2026 FDA Warning Letter to Baxter International identifies deficiencies in cross-contamination prevention, aseptic processing, environmental monitoring and quality investigations. The findings highlight why a documented Contamination Control Strategy (CCS) must be supported by evidence that manufacturing processes remain under control.
The US Food and Drug Administration (FDA) issued a Warning Letter to Baxter International on 29 September 2026 following an inspection of Baxter Healthcare Corporation’s manufacturing facility in Round Lake, Illinois, from 23 March to 2 April 2026. The FDA identified significant violations of current good manufacturing practice (CGMP) requirements for finished pharmaceuticals under 21 CFR Parts 210 and 211. It concluded that the facility’s manufacturing methods, facilities and controls did not conform to CGMP requirements, rendering the affected drug products adulterated under Section 501(a)(2)(B) of the Federal Food, Drug, and Cosmetic Act.
The agency also found that Baxter’s response to the inspection observations, submitted on 23 April 2026, did not adequately address the identified deficiencies. In its quarterly filing for the period ending 30 September 2026, Baxter subsequently disclosed that it had responded to the Warning Letter in October and was continuing to implement corrective and preventive actions (CAPA). This company disclosure does not constitute confirmation by the FDA that the response or remediation measures are adequate. The FDA Warning Letter details the observations and the corrective actions requested from the company.
Beyond individual procedures: a systemic compliance problem
The observations covered several interconnected areas of sterile manufacturing. The FDA identified deficiencies in the separation of beta-lactam products, environmental monitoring, validation of decontamination processes, aseptic process simulation, Grade A monitoring, airflow visualisation, manual visual inspection and the investigation of deviations.
Taken together, these findings illustrate a central challenge in contamination control: the presence of individual technical measures and written procedures does not, by itself, demonstrate that contamination risks are adequately managed. Effective control depends on whether facility design, process validation, monitoring data, investigations and corrective and preventive actions (CAPA) work together to maintain a demonstrable state of control.
The Warning Letter also identifies weaknesses extending beyond the Round Lake facility. The FDA refers to similar CGMP violations at other sites within Baxter’s manufacturing network and calls for a comprehensive assessment of the company’s global manufacturing operations. This makes management oversight and the effectiveness of the wider quality system central elements of the case.
Key FDA Findings
1. Inadequate separation of beta-lactam products
The FDA found that Baxter had not ensured complete and comprehensive separation between cephalosporin manufacturing and other production areas within the shared facility. Although the building sections were separated by a physical wall and had dedicated heating, ventilation and air-conditioning (HVAC) units, environmental monitoring had detected cephalosporins in multiple areas outside the designated beta-lactam production section since January 2025.
The agency also identified shortcomings in sampling locations and frequency, the qualification of swab sampling methods, and the handling of samples beyond established solution-stability periods. Multiple failing environmental samples had been discarded before further analysis.
The FDA considered the existing separation measures insufficient because they did not adequately prevent potential contamination transfer through air, personnel, materials, equipment or other interactions between production areas. The agency requested an independent assessment of facility separation, a comprehensive contamination risk assessment and a remediation plan supported by defined timelines.
2. Deficiencies in aseptic processing and validation
The FDA identified recurring recoveries of biological indicators during requalification of decontamination processes, including positive results across multiple production lines. The agency found that acceptance criteria permitted repeated recoveries without adequate investigation and that the company had not sufficiently evaluated the variability of the process or its potential impact on products.
Aseptic process simulations, commonly known as media fills, also failed to adequately represent actual production conditions. Records showed that certain interventions occurred more frequently during routine manufacturing than they were simulated during media fills.
The FDA stated that simulations that do not reflect contamination risk factors and actual production conditions cannot reliably demonstrate the state of process control or assurance of sterility.
3. Environmental monitoring and airflow visualisation
The agency found insufficient monitoring of critical processing areas, including inadequate viable-air sampling and a lack of microbiological surface monitoring in certain areas. Airflow visualisation studies were missing for some production lines or did not adequately assess worst-case operating conditions. The studies that were available revealed substantial turbulence in areas containing processing equipment.
The FDA also noted a history of sporadic positive sterility tests dating back to 2013 and persistent particulate contamination issues. The agency requested a comprehensive assessment of aseptic processing design, equipment, cleanrooms, airflow, personnel and material movements, together with improved monitoring and remediation plans covering both interim risk mitigation and long-term corrective measures.
4. Inadequate manual visual inspection
The FDA found that Baxter’s manual visual inspection programme did not reliably demonstrate the ability of operators to detect visible particles in sterile injectable products. Observations included inadequate operator qualification, oversized particles in challenge kits and inspection speeds that did not align with the standards referenced in company procedures.
The agency also cited particulate contamination complaints and a May 2025 investigation in which foreign material consistent with cardboard was identified in a returned cefazolin sample. FDA noted that Baxter agreed to a voluntary recall of one lot of cefazolin and various other products in distribution following discussions with the agency in July 2026.
The recall concerned cefazolin in dextrose injection, lot LD175708. Baxter’s recall announcement identified cardboard-like particulate matter in the solution. The FDA requested an independent assessment of the visual inspection programme, including qualification, acceptance and rejection criteria, equipment, validation, complaint data and the evaluation of unexpired batches distributed from the facility.
5. Weak investigations and CAPA effectiveness
The Warning Letter identified inadequate investigations into container-closure integrity failures, sterility test results and other critical deviations.
In one case involving cefazolin, the company invalidated a sterility test result and released the batch without an adequate investigation to establish the root cause and assess the potential impact on product quality. The FDA also noted that leak defects had increased over several years while the number of non-conformance reports had decreased.
The agency requested comprehensive assessments of the investigation and CAPA systems, including root cause analysis, trend evaluation, effectiveness checks and stronger quality-unit oversight.
The FDA additionally questioned the independence, authority and resources of the quality unit and requested a comprehensive review of management oversight. It recommended that Baxter engage a qualified CGMP consultant to conduct a six-system audit covering the quality system, facilities and equipment, materials, production, packaging and labelling, and laboratory controls.
Why the findings matter for ATMP manufacturing
The Baxter case is also relevant to manufacturers of advanced therapy medicinal products (ATMPs), including cell and gene therapies. Many such products cannot undergo terminal sterilisation, making sterility assurance particularly dependent on aseptic processing, appropriately designed facilities, controlled transfers, validated processes and effective environmental monitoring.
For these operations, manual interventions and interactions between personnel, materials and equipment can introduce contamination risks. A risk-based approach should therefore identify and assess interventions, determine whether they can be eliminated or replaced by closed-system operations, and ensure that process simulations reflect the most challenging conditions encountered in routine manufacturing.
The Warning Letter does not establish ATMP-specific findings or requirements. Its relevance to ATMP manufacturers lies in the broader manufacturing principles it illustrates: contamination risks must be assessed across the complete process, and the effectiveness of technical and organisational controls must be demonstrated with appropriate evidence.
From documented procedures to demonstrable control
A robust Contamination Control Strategy (CСS), where applicable, should connect facility and equipment design, qualification, environmental monitoring trends, aseptic process simulations, airflow visualisation, deviation investigations and CAPA effectiveness.
For US manufacturers, the applicable CGMP obligations arise from the relevant FDA regulations and guidance. The CCS terminology is also prominent in the EU GMP framework, particularly Annex 1. Although the regulatory terminology differs, the underlying objective is to establish and maintain effective, risk-based contamination controls.
The purpose is not simply to demonstrate that procedures exist, but to show how the combined controls prevent, detect and address contamination risks throughout manufacturing.
For ATMP facilities, a useful internal assessment is whether the quality unit can trace the relationship between a specific operator intervention, the relevant airflow study, environmental monitoring locations, the corresponding process simulation, the risk assessment and the effectiveness of corrective actions. If these elements are documented independently but are not supported by a coherent risk assessment and relevant data, the existence of a formal CCS may not be sufficient to demonstrate effective contamination control.
The Baxter Warning Letter reinforces the importance of evidence-based contamination control in sterile manufacturing. The central compliance challenge is to demonstrate that identified risks are understood, monitoring systems provide meaningful early warning, investigations establish scientifically justified root causes, and corrective actions deliver sustained improvements.
The case also illustrates that contamination control cannot be separated from the wider pharmaceutical quality system. Facility design, process capability, laboratory controls, documentation, management oversight and CAPA effectiveness must operate as an integrated system to support sustained CGMP compliance.